Medical device manufacturing and sales industry, manufacturing industry, manufacturing and sales approval and certification
Overview of the Pharmaceutical and Medical Device Act
Updated: July 2024, 9
Due to the revision of the Pharmaceutical Affairs Law to the Pharmaceutical and Medical Device Law, in order to ensure the safe and prompt provision of drugs, medical devices, etc., the obligation to submit package inserts has been created, and the scope of certification by registered certification bodies for medical devices has been expanded, etc. Safety measures were strengthened, and regulations were established based on the characteristics of medical devices and in vitro diagnostic drugs.
Main revisions regarding medical devices
Changes related to medical device manufacturing industry
(1) Manufacturing industry changes from permit/certification system to registration system
Previously, domestic manufacturers were subject to a permit system, and foreign manufacturers were subject to certification, but both have been changed to a registration system and requirements have been simplified. The previous manufacturing classification has disappeared, and the scope (target) of manufacturing industry registration has changed.
Before revision
After revision
Permission/registration etc.
Domestic: Permit, Foreign: Certification
Domestic/Foreign: Registration
Valid period
5 years
5 years
Permit/registration holder
Domestic: prefecture, foreign: country
Domestic: prefecture, foreign: country
Manufacturing category
General, sterile, biological, packaging, etc.
None
Requirements for permission/registration, etc.
Disqualification requirements
Disqualification requirements
Structural equipment requirements (By category: general, sterile, biological, packaging, etc.)
None (required items are stipulated in the new QMS ministerial ordinance)
Responsible engineer
必要
必要
The classification of general, packaging, etc. has disappeared, and the Regulations on Structures and Facilities for Pharmacies, which was a license requirement under the Pharmaceutical Affairs Law, are no longer applied.
(2) Changes in manufacturing process that require registration
Facilities that perform "design," which was previously not subject to permission, are now subject to registration, and "programs" and "recording media on which programs are recorded" also need to be registered as medical devices.
Manufacturing process
①Medical equipment (Other than ②③④)
②General medical equipment
③Single program
④Single program recording medium
Design
○
×
○
○
Main assembly
○
○
×
×
sterile
○
○
×
×
Storage of final products in Japan
○
○
×
○
Changes to the definition of medical devices
"Programs" themselves, which were not subject to regulation under the Pharmaceutical Affairs Law, are now subject to regulations based on international consistency, and "programs" and "recording media on which they are recorded" have been added to the definition of medical devices.
The scope of programs applicable to medical devices is as follows:
Program
1. Disease diagnosis program
2. Disease treatment programs
3. Disease prevention programs
Recording medium that recorded the program
1. Recording medium that records disease diagnosis programs
2. Recording medium that records disease treatment programs
3. Recording media that records disease prevention programs
Review of licensing requirements and compliance matters related to medical device manufacturing and sales business (QMS related, etc.)
Reorganization of QMS system
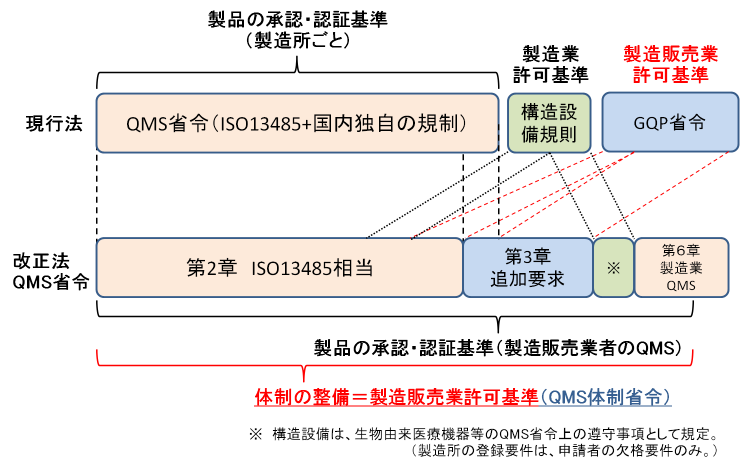
Compliance investigations with standards for manufacturing control and quality control of medical devices will be conducted for each quality control supervision system, including the product's marketing authorization holder and all registered manufacturing facilities, instead of the previous "inspection for each manufacturing facility." A new regulatory system will be applied to conduct investigations.
(1) The QMS Ministerial Ordinance has been revised and now applies to manufacturing and sales businesses.
Until now, medical device manufacturers and distributors were required to keep records based on GQP/GVP procedure manuals, but GQP has been changed to QMS, and related manufacturers have been integrated into the Quality Management Supervision System Standards (Quality Manual). ) is required.
(2) Compliance with the QMS System Ministerial Ordinance has been added as a license requirement for manufacturing and sales of medical devices, etc., in place of the GQP Ministerial Ordinance.
The Ministerial Ordinance Concerning Standards for Systems for Manufacturing Control or Quality Control of Medical Devices or In-Vitro Diagnostic Reagents (Ministry of Health, Labor and Welfare Ordinance No. 26 of August 8, 8) applies to manufacturers and distributors who handle only general medical devices. This also applies to businesses.
(3) Rationalization of surveys by product group
Product Group Ministerial Ordinance (Ministerial Ordinance that stipulates the classification of medical devices or in vitro diagnostic drugs as stipulated in Article 23-XNUMX-XNUMX, Paragraph XNUMX, Item XNUMX of the Act on Ensuring the Quality, Efficacy, and Safety of Pharmaceuticals, Medical Devices, etc.) ) has been enacted, and investigations are no longer required for products of the same product group and from the same registered manufacturing facility.
Changes regarding the certification system for medical devices
(1) The scope of third-party certification by registered certification bodies has been expanded. Not only controlled medical devices, but also "highly controlled medical devices specified by the Minister of Health, Labor and Welfare based on standards" are now subject to certification by private registered certification bodies.
(2) Under the Pharmaceutical Affairs Law, succession of the status of the holder was allowed for approved items, but under the Pharmaceutical and Medical Device Law, succession is also possible for certified items.
Regulations on sales and rental of medical devices
Not only rentals for which compensation is obtained, but also repeated rentals without compensation are subject to permission or notification, and the name has been changed from rental business to lending business.
For consultation regarding permits and licenses regarding medical devices, please contact Support Administrative Scrivener Corporation.
At Support Administrative Scrivener Corporation, we provide application support and consulting regarding the Pharmaceutical and Medical Device Act, from those newly entering the medical device industry to existing marketing distributors, manufacturers, and distributors.
Applications for medical devices are highly specialized and can be said to be one of the fields in which there are few administrative scriveners who can handle them.
We provide legal services related to medical devices based on the know-how and track record that we can accumulate because we are administrative scriveners who handle applications and negotiations with administrative agencies on behalf of companies on a daily basis.
Our staff has a track record of applying in all prefectures across the country. Please contact us by all means.